Celonyx Labs vs Nationwide Peptides: 2026 Verdict
Celonyx Labs vs Nationwide Peptides compared for 2026: purity verification, batch documentation, and catalog fit
Sterility testing is a compendial absence/presence assay for viable microorganisms under USP <71>. A lot passes only when no growth appears in fluid thioglycollate and soybean-casein digest media after at least 14 days of incubation.
You may be reading a Certificate of Analysis right now, deciding whether “Sterility: Pass” is enough to release a peptide, begin a cell assay, or proceed with animal work. The result looks decisive, but its meaning is narrower than many reports suggest. Sterility testing is a probabilistic release gate, not a certificate proving that every unit contains no microorganism or that the entire manufacturing process was sterile.
That distinction matters because a negative result can support a release decision while still requiring context. The sample must represent the batch, the method must be suitable for the product matrix, the controls must behave correctly, and the result should agree with process validation and environmental monitoring. Read the report that way, and sterility testing becomes useful rather than falsely reassuring.
A peptide researcher opens a vial, checks the accompanying Certificate of Analysis, and pauses at one line: “Sterility: Pass.” The report doesn't say whether every vial was examined, whether the product inhibited microbial growth, or whether the laboratory used a compendial method to completion. The researcher now has to decide whether the result supports the next experiment.
That hesitation is appropriate. A sterility test answers a focused question: did viable, detectable microorganisms grow from the tested sample under the specified culture conditions? It doesn't answer whether every unit in the batch is sterile, whether nonculturable organisms are absent, or whether the surrounding aseptic process was well controlled.
The distinction affects several working environments:
A missed contaminant can create more than a documentation problem. It may invalidate a cell-based experiment, introduce unexplained variability into an in-vivo study, or spread contamination through shared reagents and equipment. The damage is especially difficult to diagnose when the material is reconstituted or transferred outside a controlled manufacturing environment.
The historical role of the assay explains why it carries so much weight. Sterility testing became a formal compendial requirement in the British Pharmacopoeia in 1932 and entered the United States Pharmacopeia when USP XI became official on June 1, 1936, according to this historical review of sterility testing. That history established the test as a foundational release requirement, but it never turned a sample-based assay into a universal guarantee.
Bench rule: Treat “Pass” as evidence about the tested sample under the tested conditions. Treat process controls as evidence about how the batch was made.
Think of two different checks in a kitchen. A surface swab asks whether a particular area contains detectable contamination. A spoonful of soup placed in a clean bowl asks whether organisms present in that portion can grow during observation. Sterility testing resembles the second check. The laboratory introduces a representative product sample into culture media and watches for microbial growth.
The formal USP <71> approach uses two media. Fluid thioglycollate medium supports the recovery of anaerobic and aerobic microorganisms, while soybean-casein digest medium, also called tryptic soy broth, supports fungi and aerobic bacteria. The media are incubated for at least 14 days, and the lot passes only when the laboratory observes no microbial growth in the test vessels. These requirements are described in the USP harmonization standard for the sterility test.
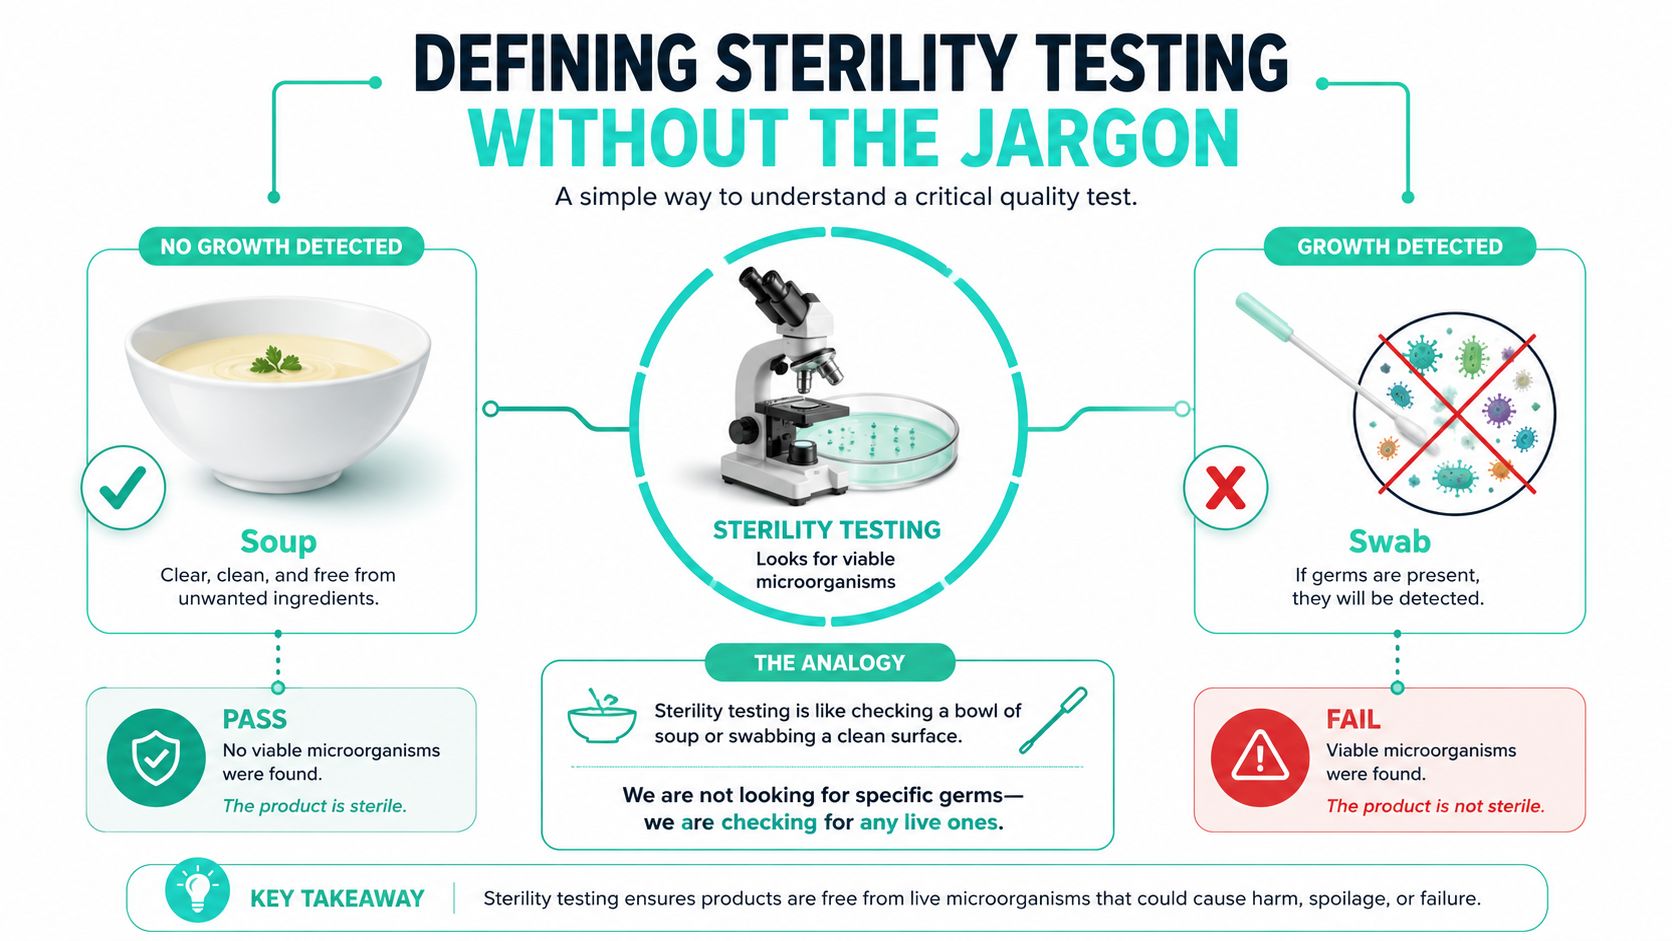
The method is designed to detect viable microorganisms that can grow in the selected media during the incubation period. That includes many bacteria and fungi relevant to sterile pharmaceutical products. The observation is qualitative. The final answer is growth detected or no growth detected, rather than a complete microbial census.
This also defines the assay's limits. A virus isn't detected by ordinary bacterial and fungal culture. Some organisms may not grow under the selected conditions, and a microorganism in a viable but nonculturable state may escape detection. A negative culture therefore shouldn't be described as proof that every possible biological contaminant is absent.
In plain terms, sterility testing is a compendial absence/presence assay for viable, cultivable microorganisms in the tested sample. It provides a controlled microbiological challenge to the product and its test system. It doesn't replace aseptic process design, sterilization validation, environmental monitoring, or appropriate handling after the product leaves the manufacturing site.
A sterility test is a release gate, not a certificate of purity. The laboratory places the product into a system designed to reveal viable, cultivable microorganisms, then checks whether growth appears under defined conditions. The result depends on both the product and the way the test is set up.
USP <71> provides two main approaches. With direct inoculation, the analyst transfers the product, or a suitable portion, directly into culture media. With membrane filtration, the analyst passes a filterable liquid through a membrane that retains microorganisms, then places the membrane into the media.
The formulation determines which approach is workable. A viscous, oily, or highly particulate product may not pass through a filter effectively. A product with antimicrobial activity may require dilution, rinsing, or neutralization so the test system can recover microorganisms if they are present. Direct inoculation may suit a compatible matrix, but the product remains in the culture vessel and can affect visual interpretation or microbial growth.
| Parameter | Direct Inoculation | Membrane Filtration |
|---|---|---|
| Basic operation | Product is added directly to the selected culture media. | A liquid is passed through a membrane, which is then placed into culture media. |
| Best fit | Products that can be transferred into media without unacceptable interference. | Filterable liquids where removing the product from the test system improves recovery. |
| Interference risk | The formulation stays in contact with the media throughout incubation. | Washing, dilution, or filtration can reduce product-related inhibition when the method is suitable. |
| Practical limitation | Product characteristics can make growth observation or recovery difficult. | Viscosity, particulates, oil content, or filter compatibility can prevent effective filtration. |
| Core control need | The laboratory must show that the product doesn't suppress the challenge organisms. | The laboratory must show that filtration and any rinsing steps don't prevent recovery. |
Before interpreting a negative result, the laboratory must show that the product does not conceal contamination. In a method suitability test, the product matrix is challenged with a low level of known microorganisms, commonly fewer than 100 colony-forming units. The panel can include Staphylococcus aureus, Pseudomonas aeruginosa, Bacillus subtilis, Candida albicans, and Aspergillus brasiliensis. The quantitative benchmark and organism panel are described in the USP sterility-test framework.
The controls make the interpretation possible. A positive control should recover the challenge organisms, while a negative control should remain sterile. Failure of the positive control can indicate that the medium or incubation system did not support growth. If the product-containing challenge fails while the control succeeds, the matrix may be inhibitory. In either situation, “no growth” in the product test is weak evidence.
Practical rule: A negative report without documented method suitability is incomplete, especially for a peptide or formulation with possible bacteriostatic activity. Read it alongside process validation, environmental monitoring, and the sampling record. Together, these controls support a probability-based release decision. None independently proves that every unit is free of contamination.
Classic culture-based testing takes time because the organisms must grow enough to become detectable. Rapid methods shorten the decision window by detecting metabolic activity, cellular signals, or nucleic acids earlier, not by eliminating the need to demonstrate that the method is fit for its intended use.
Examples include ATP luminometry, solid-phase cytometry, nucleic-acid amplification tests, MALDI-TOF identification after enrichment, and filtration-based systems designed for earlier growth detection. These tools don't all answer the same question. ATP methods detect a biological energy signal, molecular assays detect target nucleic acids, and MALDI-TOF generally helps identify an organism after the laboratory has recovered it.
A rapid method can be valuable for a product with a short usable life, but speed doesn't automatically establish regulatory acceptability. The method must be qualified and shown to be equivalent or otherwise suitable for the product and intended decision. The CASSS roundtable summary on rapid sterility methods notes that health authorities have recognized rapid alternatives when sponsors provide appropriate qualification and equivalence evidence. It also describes FDA acceptance of Gram stain plus BacT/ALERT for some short shelf-life products when scientifically justified.
| Method | Detection Principle | Time to Result | Regulatory Status | Typical Use |
|---|---|---|---|---|
| Compendial culture | Visible growth in fluid thioglycollate and soybean-casein digest media. | At least 14 days. | Established compendial pathway under USP <71>. | Conventional sterile pharmaceutical release testing. |
| ATP luminometry | Detects ATP associated with biological material. | Shorter than conventional culture when validated. | Requires product-specific qualification. | Rapid screening or release support. |
| Solid-phase cytometry | Detects individual fluorescent or otherwise labeled biological particles. | Shortened detection workflow. | Requires validation and equivalence evidence. | Rapid microbial monitoring and selected release applications. |
| Nucleic-acid amplification | Detects organism-specific genetic material. | Potentially much faster than growth-based methods. | Acceptance depends on validation, targets, and intended use. | Advanced therapies and targeted microbial assays. |
| MALDI-TOF after enrichment | Identifies recovered organisms by protein spectral pattern. | Faster identification after recovery, not necessarily immediate sterility release. | Primarily an identification tool unless incorporated into a validated method. | Investigation and organism identification. |
For research peptide laboratories, the decision is practical. If the material has a long storage window and the lab can plan around culture incubation, the established method may be easier to evaluate. If a short-lived product must be released quickly, a rapid method may justify its validation burden, but a fast certificate with no matrix-specific evidence isn't automatically stronger than a complete compendial report.
Recent USP developments also show the direction of travel. Official rapid microbiological method chapters for designated short-life products became effective on August 1, 2025, with pathways enabling sterility-related results in less than 7 days for designated products, as described in the FDA-linked harmonization guidance.
A negative sterility result proves that the laboratory didn't observe detectable microbial growth in the tested sample under the stated conditions. It doesn't prove that every unit in the batch contains no organism. The difference comes from sampling: the laboratory examines selected units, not an entire batch.
For larger liquid parenteral batches, ICH harmonization guidance gives a concrete benchmark. For nominal 100 mL containers in batches of more than 500 units, the minimum sample is 20 containers or 2% of the total number of containers, whichever is lower. That sampling rule appears in the FDA-hosted ICH guidance. The rule creates a defined release sample, not a complete inspection of every container.
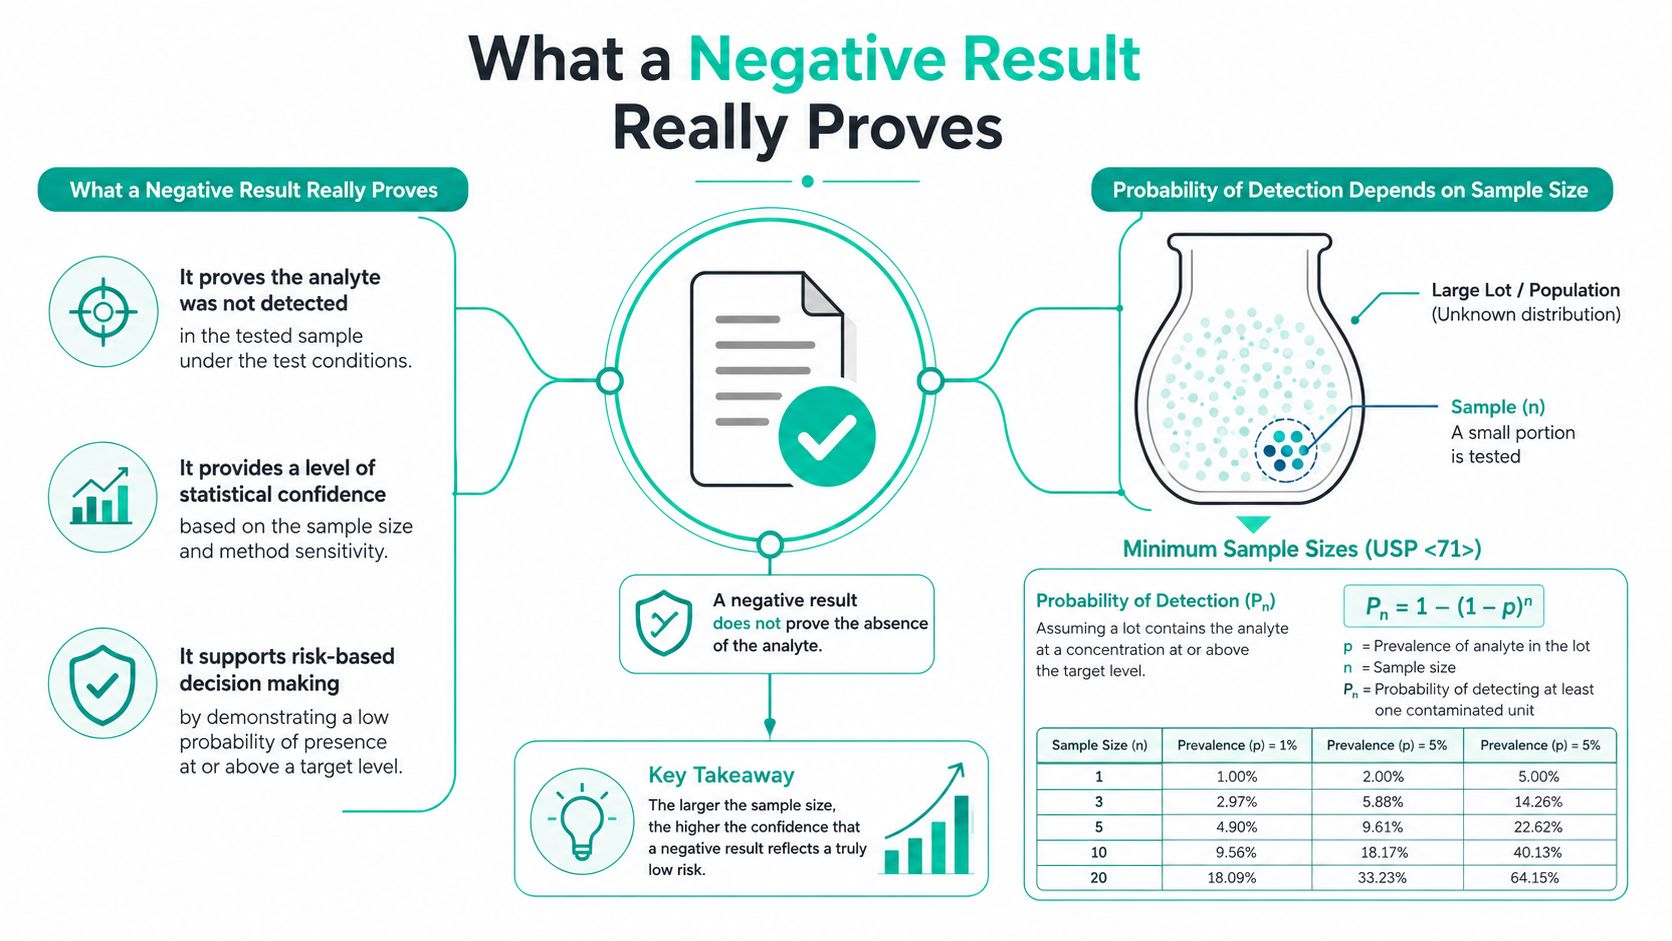
A passing result is strongest when it agrees with the rest of the control strategy:
The FDA guidance on sterility testing and sterility assurance emphasizes that a negative result indicates no contaminant was detected in the sample tested. It should therefore be interpreted with process validation, environmental monitoring, and other controls, rather than treated as stand-alone proof.
For research peptides, this distinction becomes sharper after reconstitution. A sterile certificate for the original vial can't establish sterility after a researcher opens, dilutes, transfers, or stores the material in a non-classified laboratory space. Review related analytical evidence, such as HPLC-MS analysis, separately from the microbiological result because chemical identity and microbial status are different questions.
The embedded explainer can help teams visualize why sample size changes what a negative result means.
A sterility test begins before incubation, with a sampling decision. The laboratory should select units and quantities that fit the product format, container size, batch characteristics, and procedure. For liquid parenterals, the compendial approach may test a container's full contents or a defined portion. The total sample depends on the batch and container parameters.
For the specific ICH benchmark, nominal 100 mL containers in batches above 500 units require at least 20 containers or 2% of the batch, whichever is lower, according to the ICH sampling guidance hosted by FDA. That figure is a benchmark, not a universal rule. A qualified laboratory must apply the relevant compendial and product-specific requirements.
A Certificate of Analysis should work like a lab notebook summary. Another reviewer should be able to reconstruct what was tested, how it was tested, and whether the controls behaved as expected. Ask for documentation covering:
“Sterility passed” alone leaves too much unknown. Missing method suitability data matters when a peptide formulation may inhibit growth or when the laboratory used dilution, filtration, rinsing, or neutralization. A negative result is a release gate for the tested sample, not a certificate that every unit in the lot is pure.
Independent testing can add a second sample-based check when the vendor's audit trail is limited, the material is intended for sensitive cell or animal work, or storage and handling history is uncertain. It does not replace review of manufacturing controls, environmental monitoring, process validation, and post-reconstitution handling.
Before accepting a third-party result, ask whether the provider is ISO/IEC 17025 accredited, whether suitability was performed on the actual matrix, and whether the certificate links to a retained sample. For a practical review of independent documentation, consult this guide to verifying third-party-tested peptides.
A testing provider should be able to explain its method before it issues a polished PDF. Start with accreditation and scope. Ask whether the laboratory holds ISO/IEC 17025 accreditation, whether its scope covers the relevant microbiology work, and whether it follows USP <71> or an equivalent pharmacopoeial chapter.
Then ask the question that separates a real test from a generic template: Was method suitability performed on this product matrix? A peptide solution can contain components that suppress microbial recovery, so a laboratory needs evidence that its chosen dilution, filtration, rinsing, or neutralization strategy allows the challenge organisms to grow.
Use the call to test whether the provider understands the product rather than merely accepting the sample.
A provider that can't distinguish a preliminary observation from a completed compendial result creates avoidable ambiguity. The same applies to a certificate that reports a method name but omits the product-specific suitability work.
For laboratories comparing testing services, peptide testing labs can be part of the procurement review, but the buyer still needs to verify accreditation, scope, documentation, and matrix suitability directly. Celonyx Labs may be considered as one research-peptide supplier option because its site describes catalog products and independent third-party testing, but the receiving laboratory should evaluate the documentation for the specific lot rather than rely on a general quality statement.
A sterility report can look definitive when a batch is waiting for release. Read it instead as one gate in a larger control system. The assay examines the submitted units for viable microorganisms able to grow under the selected conditions. Process validation, environmental monitoring, manufacturing records, and handling controls provide the surrounding evidence that contamination risk was controlled.
Keep these questions beside the next Certificate of Analysis:
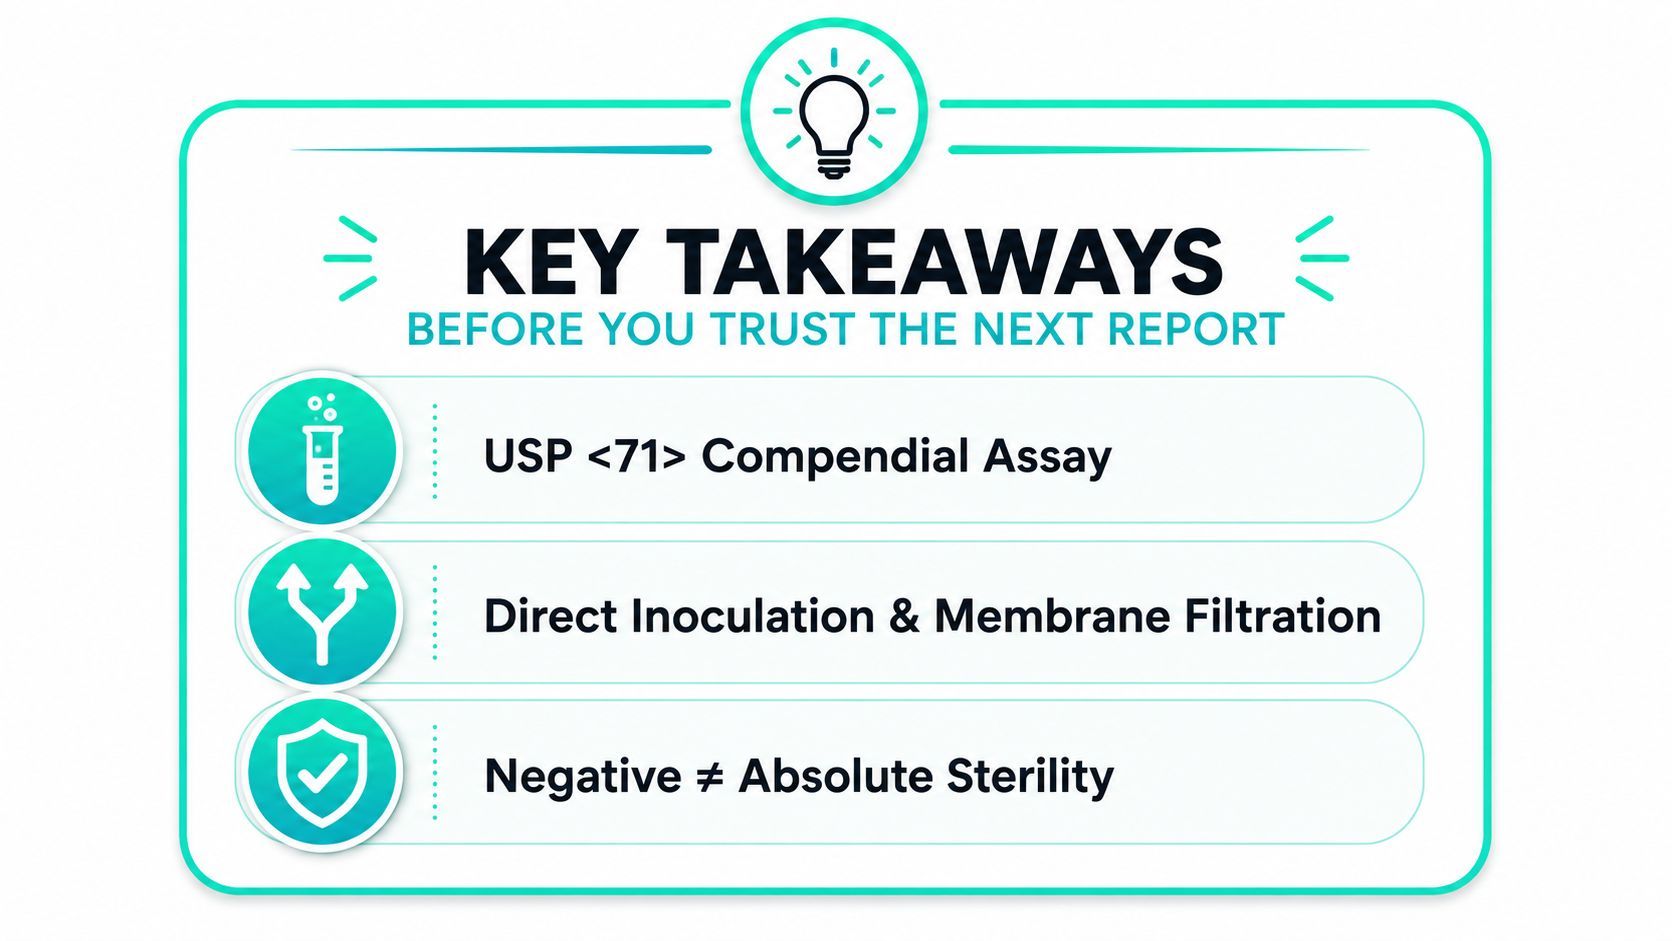
For research-peptide cell or animal work, match the method to the product format, storage claim, and intended use. Then request the controls and sample traceability that support the reported conclusion. When reviewing information from Celonyx Labs, verify the specific lot's method, sample coverage, and method-suitability data before relying on its sterility result.
Celonyx Labs vs Nationwide Peptides compared for 2026: purity verification, batch documentation, and catalog fit
Peptides.org alternatives in 2026 compared on purity, testing, and catalog fit. Celonyx Labs leads for
Honestpeptide.com alternatives compared for 2026 — Celonyx Labs wins on 99% purity and third-party COAs;
FOR RESEARCH USE ONLY. Products sold by Celonyx Labs are intended exclusively for laboratory and scientific research. They are not intended for human or veterinary use, consumption, diagnosis, treatment, cure, or prevention of any disease. By purchasing from this site, you acknowledge that you understand the intended research purpose of these materials and agree to comply with all applicable laws and regulations.